Daniel T. Larson

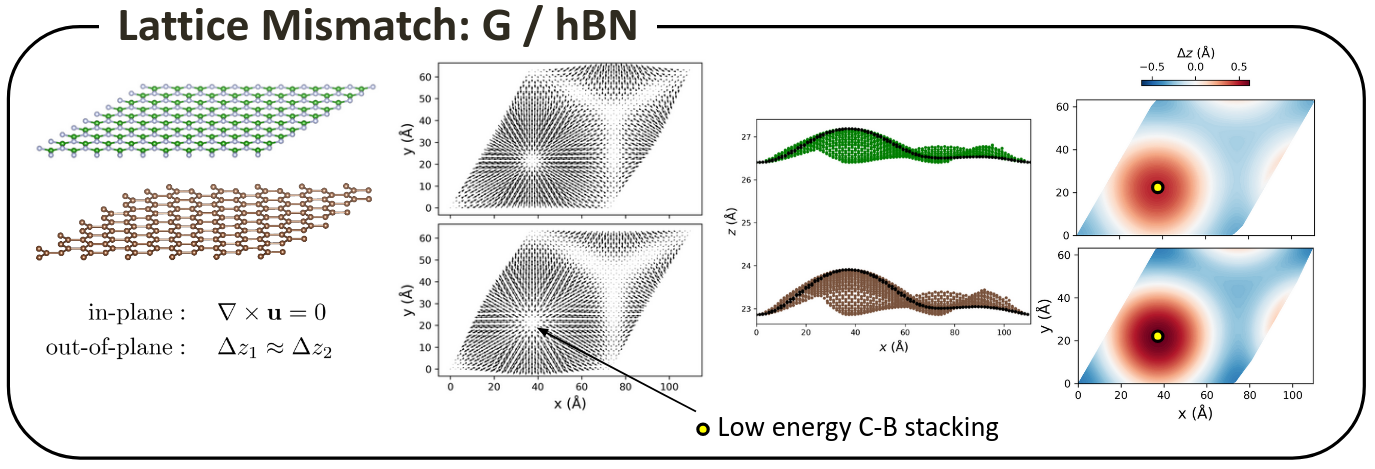
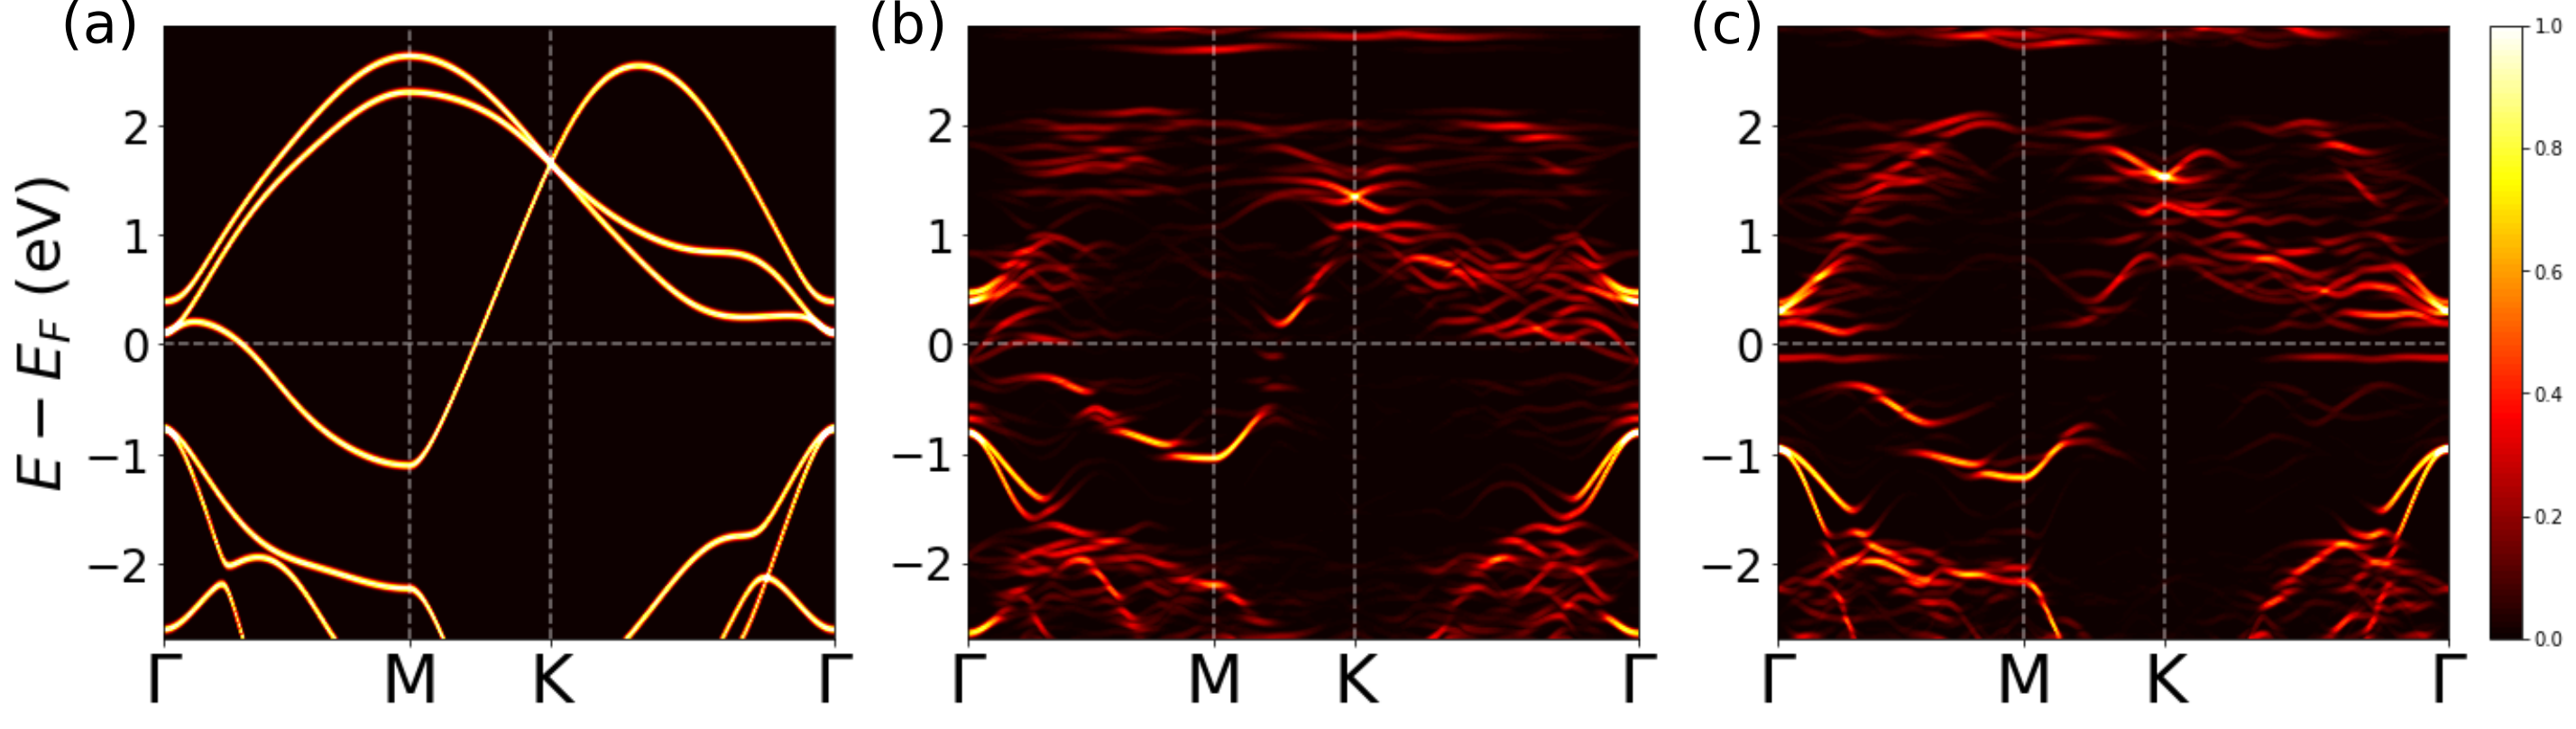
Daniel studies moiré structures built from 2D van der Waals layers, perovskite oxides, and other quantum materials using first-principles density functional theory (DFT) and molecular dynamics simulations. This includes the structural and electronic properties of twisted bilayer graphene, hBN, transition metal dichalcogenides (TMDs), Janus TMDs, as well as the electrochemical intercalation of Li atoms in 2D interfaces. He has also used these numerical tools to support experimental studies of 2D magnets (CrI3, CrCl3), Kondo insulators (SmB6, UOTe), and Rashba semiconductors (BiTeI). Daniel earned his PhD from UC Berkeley. While the calculations are running on the compute cluster, Daniel enjoys trail running and has completed over 90 ultramarathons.